Have you ever heard of a rare condition called Pompe disease? You might find this name completely new and unfamiliar. It is a very rare condition that does not happen to everyone. However, learning about it is very important because it is passed down through your family genes. When you have this condition, a specific type of sugar called glycogen builds up inside your cells. Today, let us talk simply about what Pompe disease is, why it happens, what signs to look for, and what treatments doctors use.
Living with or caring for someone with a rare medical condition can feel overwhelming at first. Understanding the basics helps you know what to expect and when to talk to your doctor. Let us break down everything you need to know about Pompe disease in clear, everyday language.
Key Takeaways About Pompe Disease.
- Genetic Cause: Pompe disease is an inherited condition caused by changes in your GAA gene.
- Enzyme Shortage: Your body lacks enough acid alpha-glucosidase enzyme to break down stored sugar.
- Two Main Types: It can start in infancy or appear later in childhood or adulthood.
- Muscle Impact: The buildup of sugar mainly weakens your skeletal muscles and heart muscle.
- Effective Treatment: Enzyme replacement therapy helps manage symptoms and improves your daily life.
Know your body mass index (BMI) and weight classification
What is Pompe Disease?
Simply put, Pompe disease belongs to a group of conditions called glycogen storage disorders. Inside your body, you have an important enzyme called acid alpha-glucosidase, also known as GAA. This enzyme helps break down complex sugar molecules called glycogen so your body can turn them into energy. If you have Pompe disease, your body does not make enough of this GAA enzyme, or it does not work correctly.
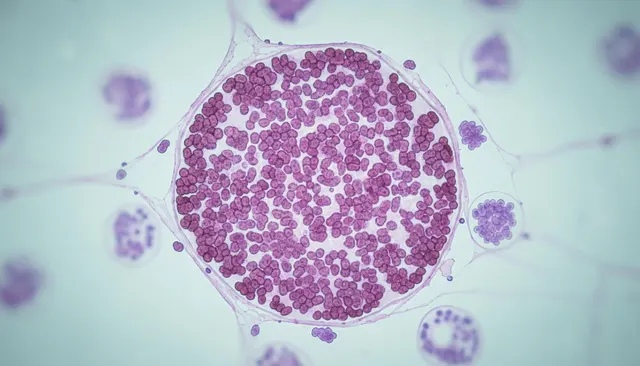
When this enzyme is missing or low, glycogen cannot break down properly. Instead, it builds up inside tiny compartments in your cells called lysosomes. Think of lysosomes like tiny recycling bins inside your cells. Because the GAA enzyme is missing, glycogen piles up inside these bins until they overflow. This buildup happens mostly in your heart muscle and skeletal muscles. Over time, this damage makes your organs and muscles weaker and less able to do their jobs.
Doctors also call Pompe disease by a few other names:
- Pompe's disease.
- Acid maltase deficiency.
- Glycogen storage disease type II, or GSD2.
Main Types of Pompe Disease.
Doctors divide Pompe disease into two main types based on when the signs start and how severe they are. Knowing the type helps your medical team choose the right care plan for you.
Infantile-Onset Pompe Disease.
This is the most severe form of Pompe disease. It happens when your body has little to no GAA enzyme activity at all. A baby might look completely healthy and show no signs right after birth. However, signs usually start within the first year of life, often around four months of age.
Babies with this type experience severe muscle weakness, an enlarged liver, and an enlarged heart known as cardiomyopathy. The condition progresses very quickly. Without prompt medical treatment, these infants may face heart failure or severe breathing problems within their first or second year of life. This makes early diagnosis absolutely critical.
Late-Onset Pompe Disease.
This type can start at any age. It might appear during late infancy without heart enlargement, during childhood, in your teenage years, or even later in adulthood. People with this type still have some working GAA enzyme, which is why it develops more slowly.
This form is usually milder than the infantile type, but severity depends on when signs first appear. Children who show signs early in childhood can still face significant health challenges. The main sign is progressive muscle weakness, which can lead to breathing difficulties. Fortunately, your heart is rarely affected in this late-onset form. Without treatment, respiratory complications can still become life-threatening over time.
How Common is Pompe Disease?
Pompe disease is a very rare genetic condition. In the United States, for example, it affects roughly one out of every 40,000 people. While exact numbers vary around the world, health experts agree that it is an uncommon condition. Newborn screening programs in many countries now test for it early so treatment can begin right away.
Symptoms of Pompe Disease.
The main sign of Pompe disease is gradual muscle weakness. This weakness mostly affects your hips, legs, shoulders, arms, and your diaphragm, which is the large muscle beneath your lungs that helps you breathe.
Infantile-Onset Pompe Symptoms.
- Floppy muscle tone, making the baby feel very limp when held.
- An enlarged heart.
- An enlarged liver.
- An unusually large tongue.
- Poor weight gain and slow growth, known as failure to thrive.
- Trouble breathing or frequent respiratory infections.
- Difficulty feeding or swallowing milk.
- Possible hearing loss.
Late-Onset Pompe Symptoms.
These signs are usually milder at first and worsen very slowly over time. However, they can still lead to major physical weakness and breathing challenges.
- Gradual weakening of the muscles in your legs and trunk.
- Difficulty walking, climbing stairs, or frequent falls.
- Achiness or pain in your muscles.
- Loss of the ability to run, jump, or exercise easily.
- Shortness of breath when you exert yourself.
- Morning headaches caused by breathing issues during sleep.
- Excessive daytime tiredness or fatigue.
- Unexplained weight loss.
- Trouble swallowing food safely.
- Irregular heart rhythms in some cases.
- Gradual hearing loss.
Causes of Pompe Disease.
Pompe disease happens because of changes, called mutations, in your GAA gene. This specific gene gives your body the instructions to make the acid alpha-glucosidase enzyme. As we discussed, this enzyme is responsible for breaking down glycogen into simple sugars.
Normally, your body converts glycogen into glucose whenever you need energy. The enzyme works inside the lysosomes, which act as the recycling centers of your cells. When your GAA gene has a mutation, your cells cannot produce enough working enzyme. Glycogen accumulates inside your lysosomes, swells the cells, and damages your muscle tissues.
This condition is inherited in an autosomal recessive pattern. This means you must inherit one altered gene from each of your parents to develop the disease. Your parents are usually carriers who do not show any signs of the illness themselves, but they both carry the hidden genetic change.
Complications of Pompe Disease.
Without treatment, infantile-onset Pompe disease can be fatal in early childhood due to heart and breathing failure. People with late-onset Pompe disease often experience increasing muscle weakness that eventually impacts their mobility. Many patients require breathing support devices or wheelchairs as the condition progresses over time.
How is Pompe Disease Diagnosed?
Your doctor will start by performing a physical exam and asking about your personal and family medical history. They may order a blood test to check levels of an enzyme called creatine kinase, which leaks out when muscles are damaged. A more specific test measures the actual activity level of your GAA enzyme in your blood or skin cells.
Your doctor may also recommend genetic testing to look for mutations in your GAA gene. Additional tests can evaluate how well your body functions:
- Pulmonary function tests: These measure how much air your lungs can hold and how well they work.
- Electromyography: This test checks the electrical activity of your muscles to see how well they respond.
- Heart tests: An electrocardiogram or echocardiogram checks your heart rhythm and structure.
- Sleep studies: These monitor your breathing patterns while you sleep.
| Diagnostic Test. | What It Measures. / Why Doctors Use It. |
|---|---|
| Enzyme Assay. | What It Measures.: GAA enzyme activity in blood. Why Doctors Use It.: To confirm low enzyme levels. |
| Genetic Testing. | What It Measures.: GAA gene mutations. Why Doctors Use It.: To identify specific gene changes. |
| Pulmonary Test. | What It Measures.: Lung capacity and airflow. Why Doctors Use It.: To check breathing strength. |
Treatments for Pompe Disease.
The main treatment for Pompe disease is enzyme replacement therapy, often called ERT. During this treatment, your doctor infuses a man-made version of the missing enzyme directly into your vein. Common medicines used include:
- Alglucosidase alfa.
- Avalglucosidase alfa.
These engineered enzymes work just like your natural ones should. This treatment helps reduce heart size, maintain normal heart function, improve muscle strength, and slow down the buildup of glycogen in your tissues.
You will also work with a team of medical specialists who help manage your specific symptoms. This care team often includes physical therapists, occupational therapists, respiratory therapists, cardiologists, and neurologists. Depending on your needs, therapy might include regular physical exercises, feeding tubes to ensure proper nutrition, or mechanical ventilators to help you breathe easily.
Researchers are also studying gene therapy in clinical trials. Gene therapy aims to replace faulty genes with healthy ones so your body can produce its own GAA enzyme. This research offers great hope for the future.
Can Pompe Disease Be Prevented?
Because Pompe disease is a genetic condition, you cannot prevent it. However, if you are pregnant or planning to have children, talking with a genetic counselor can help. They can explain your risks, discuss family inheritance patterns, and guide you through available screening tests.
Living with Pompe Disease.
Without early diagnosis and treatment, infantile-onset Pompe disease is often fatal during infancy due to heart and lung complications. People with late-onset Pompe disease usually live longer because the condition progresses more slowly. Your outlook depends greatly on when your symptoms started and how severe they are when treatment begins.
When to Seek Emergency Medical Care.
Call 911 (US/Canada), 999 (UK), or go to your nearest Emergency Department right away if you or your child experience severe shortness of breath, sudden chest pain, blue lips or skin, or sudden loss of consciousness. Early emergency intervention saves lives.
When to See a Doctor.
If you notice persistent muscle weakness, unexplained fatigue, or breathing difficulties in yourself or your child, schedule an appointment with your doctor right away. Early diagnosis and prompt treatment make a huge difference in managing symptoms and improving your overall quality of life.
Caring for Yourself or Your Child.
Caring for someone with Pompe disease involves emotional support as well as physical care. Talking with a counselor can help you process your diagnosis and cope with daily stress. Support groups also connect you with other families facing similar challenges.
Caregiver burnout is very common when managing a chronic illness. Remember that taking care of your own mental and physical health is just as important as caring for your loved one.
Questions to Ask Your Doctor.
When you visit your doctor, having a list of questions ready can help you feel more prepared:
- What specific type of Pompe disease do I or my child have?
- What treatment options are available right now?
- Which medical specialists should we add to our care team?
- How often will we need follow-up appointments and testing?
- Are there clinical trials or new therapies we should know about?
- Should other family members get genetic testing?
- What local support groups do you recommend?
💡 Take-Home Message.
Pompe disease is a rare genetic condition that causes glycogen to build up and weaken your muscles. While it presents serious challenges through infantile or late-onset forms, modern treatments like enzyme replacement therapy offer real help. Early detection, specialized medical care, and strong emotional support empower patients and families to live fuller, more comfortable lives. Always speak with your doctor if you have questions or notice unusual symptoms.
Sources and References:
1. Cleveland Clinic. Pompe Disease: Causes, Symptoms & Treatment. Available online: Cleveland Clinic Medical Reference.
2. Mayo Clinic. Pompe Disease - Symptoms and Causes. Available online: Mayo Clinic Patient Care Guide.
3. National Health Service (NHS UK). Pompe Disease Overview. Available online: NHS Health Information.
4. National Institutes of Health (NIH / MedlinePlus). Glycogen Storage Disease Type II. Available online: MedlinePlus Genetics Reference.
Medical Disclaimer: This educational guide is for informational purposes only and does not replace professional medical advice, diagnosis, or treatment. Always consult your doctor or qualified doctor with any questions regarding a medical condition.
No comments yet. Be the first to share your question or experience below.